Portfolio item number 1
Short description of portfolio item number 1
Short description of portfolio item number 1
Short description of portfolio item number 2
Published in PLoS Biol, 2019
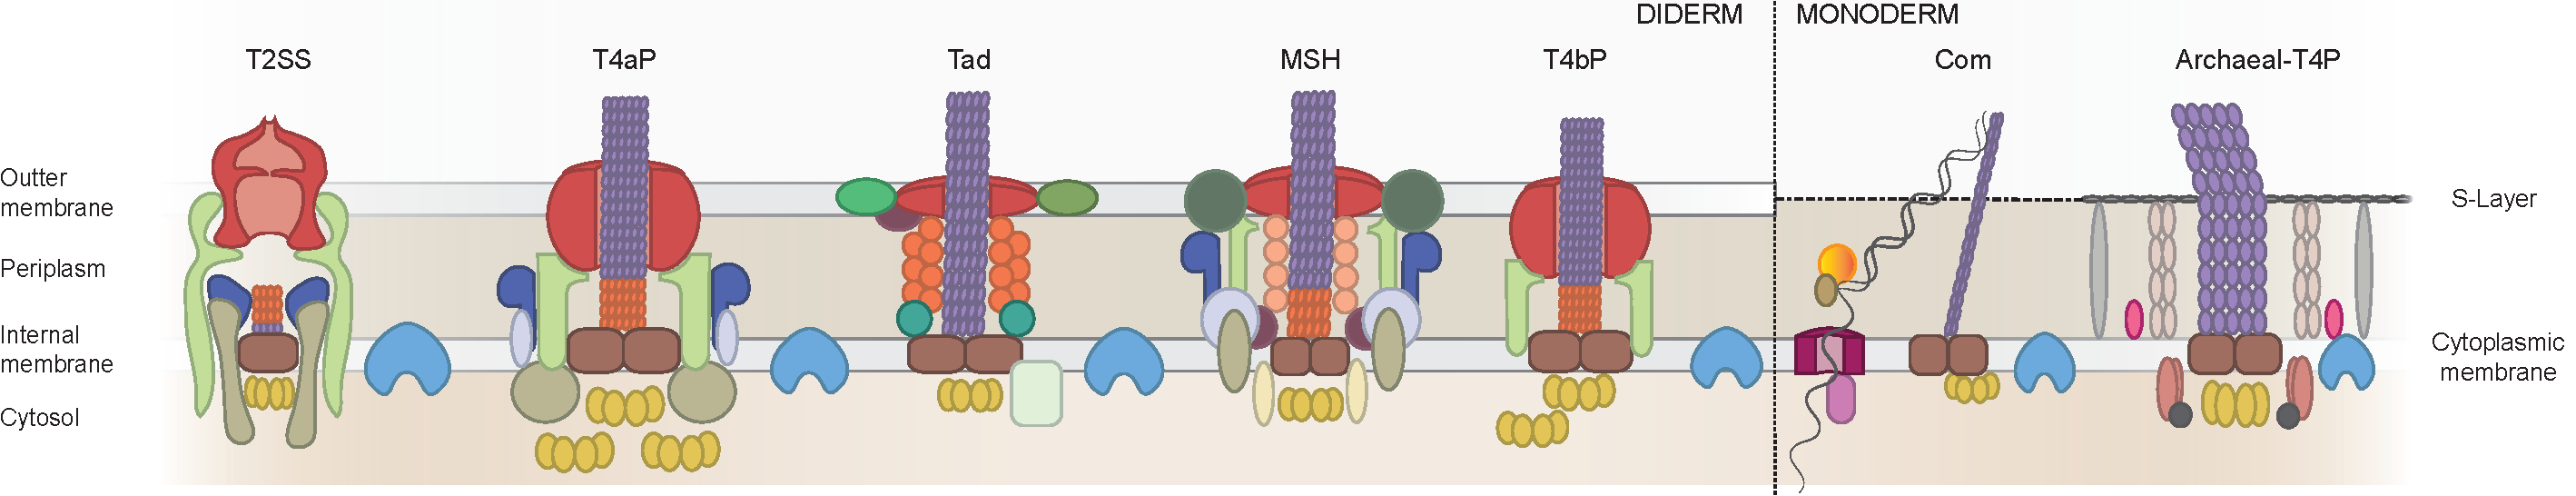
The Type IV filament (TFF) superfamily, a diverse group of membrane-associated molecular systems found in Bacteria and Archaea, has undergone significant diversification, evolving into complex “machines” responsible for various critical cellular functions. These include adhesion, protein secretion, DNA uptake, and motility. Research into the natural history of this superfamily suggests that TFF systems were likely present in the last universal common ancestor. From this ancient origin, two distinct lineages—one bacterial and one archaeal—diversified through mechanisms such as multiple gene duplications, gene fissions and deletions, and the acquisition of novel components.
Recommended citation: R. Denise, S. Abby, E. Rocha, Diversification of the type IV filament superfamily into machines for adhesion, protein secretion, DNA uptake, and motility. PLoS Biol, 2019. https://www.ncbi.nlm.nih.gov/pubmed/31323028
Published in Sorbonne Université, 2019
Process of molecular innovation require tinkering and the co-option of existing genes. But this process remains poorly understood on long evolutionary scales. During this thesis, I analyzed the evolutionary history of a large group of molecular membrane systems associated with bacteria and archaea - the superfamily of type IV filaments (TFF-SF) - that have diversified into systems involved in flagellar or contraction motility, adhesion, protein secretion, natural transformation…. I have developed tools and methods that have allowed me to identify these systems in all phyla of two of the kingdoms of life, and their phylogeny suggests that they may have been present in the last universal common ancestor. TFF-SF was then diversified by multiple gene duplications, fission of the gene from the integral membrane platform, and accretion of new components. Surprisingly, I found that Tad systems originated from the interkingdom transfer from Archaea to Bacteria of a system similar to the Epd pilus. The phylogeny and the content of ancestral systems suggest that the initial bacterial pili were involved in cellular motility and/or DNA transformation. On the other hand, specialized protein secretion systems appeared much later. All these processes of functional diversification have been accompanied by genetic rearrangements with implications for genetic regulation and horizontal gene transfer. Overall, the evolutionary history of TFF-SF reveals an impressive catalogue of the variety of molecular mechanisms involved in the origin of new functions through tinkering and co-optation of cellular machineries.
Recommended citation: Rémi Denise, Evolution of protein secretion systems in bacteria. PhD Thesis, Sorbonne Université, 2019. https://theses.fr/2019SORUS602
Published in Trends Microbiol, 2020
The article “The Evolution of Protein Secretion Systems by Co-option and Tinkering of Cellular Machineries” explores how the complex protein secretion systems in bacteria have developed. Protein secretion is crucial for various interactions, both biotic and abiotic. The evolution of these bacterial secretion systems, often described as nanomachines, primarily occurred through the “co-option” of existing cellular machinery originally used for functions like motility, conjugation, injection, or adhesion. Some secretion systems have emerged repeatedly throughout evolution, while others are unique. Their development typically involved successive cycles of gene accretion, deletion, and horizontal gene transfer.
Recommended citation: R. Denise, S. Abby, E. Rocha, The Evolution of Protein Secretion Systems by Co-option and Tinkering of Cellular Machineries. Trends Microbiol, 2020. https://www.ncbi.nlm.nih.gov/pubmed/32298615
Published in J Biol Chem, 2020
The major subunit of widespread competence (Com) pili, known as ComGC, exhibits a novel and conserved type IV pilin fold that is purely helical. This discovery stems from genomic, phylogenetic, and structural analyses of ComGC, which is the primary pilin subunit of Com pili. Type IV filaments (T4F), which are helical assemblies of type IV pilins, are filamentous nanomachines found broadly in prokaryotes and are responsible for a variety of functions. The Com pilus is a significant T4F that facilitates DNA uptake in bacteria with a single membrane (monoderms), a crucial initial step in natural transformation and an important mechanism for horizontal gene transfer.
Recommended citation: D. Sheppard, J. Berry, R. Denise, E. Rocha, S. Matthews, V. Pelicic, The major subunit of widespread competence pili exhibits a novel and conserved type IV pilin fold. J Biol Chem, 2020. https://www.jbc.org/article/S0021-9258(17)48519-X/fulltext
Published in Proc Natl Acad Sci U S A, 2021
Motor-independent retraction of type IV pili (T4P) is governed by an inherent property of the pilus filament itself, relying on its inherent instability rather than dedicated motor proteins. This mechanism appears to be a conserved feature across various Type IV Filament (T4F) systems. Research utilizing the Vibrio cholerae competence pilus as a model system has revealed key insights into this process. Mutations in the major pilin gene (PilA) that lead to enhanced motor-independent retraction likely do so by diminishing pilin-pilin interactions within the filament, resulting in less stable pili.
Recommended citation: J. Chlebek, R. Denise, L. Craig, A. Dalia, Motor-independent retraction of type IV pili is governed by an inherent property of the pilus filament. Proc Natl Acad Sci U S A, 2021. https://www.ncbi.nlm.nih.gov/pubmed/34789573
Published in Microb Genom, 2023
Pyridoxal 5’-phosphate (PLP), a cofactor derived from B6 vitamers, is crucial for the growth of all known organisms. While its synthesis and salvage pathways are understood in some model species, key components like vitamin B6 transporters remain unidentified in many organisms. A comparative genomic study predicted PLP synthesis and salvage pathways across 5840 bacterial and archaeal species with complete genomes. This research surveyed the distribution of the two known de novo biosynthesis pathways and instances of non-orthologous gene displacement. The analysis revealed that several de novo PLP pathway genes are yet to be identified in many organisms, contributing to what the researchers refer to as “pathway holes.”
Recommended citation: R. Denise, J. Babor, J. Gerlt, V. Crecy-Lagard, Pyridoxal 5,-phosphate synthesis and salvage in Bacteria and Archaea: predicting pathway variant distributions and holes. Microb Genom, 2023. https://www.ncbi.nlm.nih.gov/pubmed/36729913
Published in Peer Community Journal, 2023
MacSyFinder v2 is a major release of the Macromolecular System Finder program, designed to model and annotate complex cellular functions in microbial genomes by identifying individual genes at the molecular system level. The updated version is coded in Python 3 and features improved, rationalized code for better maintainability and more flexible system modeling. The new release introduces a more intuitive and comprehensive search engine capable of identifying optimal and sub-optimal candidate systems that adhere to model constraints. Additionally, MacSyFinder v2 includes a novel companion tool called macsydata, which facilitates the easy installation and broad distribution of models developed for MacSyFinder (macsy-models) from GitHub repositories.
Recommended citation: Bertrand Néron, Rémi Denise, Charles Coluzzi, Marie Touchon, Eduardo Rocha, Sophie Abby, MacSyFinder v2: Improved modelling and search engine to identify molecular systems in genomes. Peer Community Journal, 2023. https://peercommunityjournal.org/item/10_24072_pcjournal_250/
Published in Methods Mol Biol, 2023
The abstract for “Identification of Protein Secretion Systems in Bacterial Genomes Using MacSyFinder Version 2” describes a computational tool designed to identify and characterize protein secretion systems within bacterial genomes. MacSyFinder is a flexible tool that leverages knowledge from experimental studies to find homologous systems in genomic data. It employs predefined models, such as “TXSScan,” to identify major secretion systems in diderm bacteria, as well as related cellular appendages like pili and flagella. The process involves identifying and clustering co-localized components of secretion systems using sequence similarity searches with Hidden Markov Model (HMM) protein profiles.
Recommended citation: S. Abby, R. Denise, E. Rocha, Identification of Protein Secretion Systems in Bacterial Genomes Using MacSyFinder Version 2. Methods Mol Biol, 2023. https://www.ncbi.nlm.nih.gov/pubmed/37930518
Published in Gut Microbes, 2024
A novel lytic bacteriophage, designated phiPDS1 (φPDS1), which infects Parabacteroides distasonis, has been successfully isolated and characterized from human feces. This discovery contributes to a better understanding of the viral component of the human gut microbiome. phiPDS1 exhibits siphovirus morphology and has been classified within a newly proposed genus, Sagittacolavirus. The phage is lytic, meaning it replicates within and then lyses its host bacterial cells. Despite its lytic nature, phiPDS1 has demonstrated an ability to coexist with its host in liquid culture for several days without impacting the host’s abundance.
Recommended citation: A. Cortes-Martin, R. Denise, E. Guerin, S. Stockdale, L. Draper, R. Ross, A. Shkoporov, C. Hill, Isolation and characterization of a novel lytic Parabacteroides distasonis bacteriophage phiPDS1 from the human gut. Gut Microbes, 2024. https://www.ncbi.nlm.nih.gov/pubmed/38178369
Published in Microb Genom, 2024
“Beyond Blast” refers to a published workflow designed to assist microbiologists in more effectively extracting literature, taxonomic distributions, and gene neighborhood information for protein families. This approach aims to provide a comprehensive understanding of protein families beyond what traditional BLAST analyses offer. The workflow was developed and evaluated using a dataset of over 280 references related to the DUF34 protein family. The researchers assessed the efficiency of various databases and search tools to create a method that helps experimentalists capture the most relevant published information on protein families in the shortest time.
Recommended citation: C. Reed, R. Denise, J. Hourihan, J. Babor, M. Jaroch, M. Martinelli, G. Hutinet, V. Crecy-Lagard, Beyond blast: enabling microbiologists to better extract literature, taxonomic distributions and gene neighbourhood information for protein families. Microb Genom, 2024. https://www.ncbi.nlm.nih.gov/pubmed/38323604
Published in Phage (New Rochelle), 2025
Background: Bacteriophages are classified into genera and species based on genomic similarity, a process regulated by the International Committee on the Taxonomy of Viruses. With the rapid increase in phage genomic data there is a growing need for automated classification systems that can handle large-scale genome analyses and place phages into new or existing genera and species. Materials and Methods: We developed taxMyPhage, a tool system for the rapid automated classification of dsDNA bacteriophage genomes. The system integrates a MASH database, built from ICTV-classified phage genomes to identify closely related phages, followed by BLASTn to calculate intergenomic similarity, conforming to ICTV guidelines for genus and species classification. taxMyPhage is available as a git repository at https://github.com/amillard/tax_myPHAGE, a conda package, a pip-installable tool, and a web service at https://phagecompass.ku.dk. Results: taxMyPhage enables rapid classification of bacteriophages to the genus and species level. Benchmarking on 705 genomes pending ICTV classification showed a 96.7% accuracy at the genus level and 97.9% accuracy at the species level. The system also detected inconsistencies in current ICTV classifications, identifying cases where genera did not adhere to ICTV’s 70% average nucleotide identity (ANI) threshold for genus classification or 95% ANI for species. The command line version classified 705 genomes within 48 h, demonstrating its scalability for large datasets.
Recommended citation: A. Millard, R. Denise, M. Lestido, M.T. Thomas, D. Webster, S. D. Turner, T. Sicheritz-Pontén, taxmyPHAGE: automated taxonomy of dsDNA phage genomes at the genus and species level. Phage (New Rochelle), 2025 Mar 17;6(1):5-11. https://pubmed.ncbi.nlm.nih.gov/40351403/
Published in Cell Reports, 2025
Antibiotic use disrupts the gut microbiota, posing risks of long-term health issues and resistance. To study its impact on gut phages, we followed 22 healthy individuals 2 weeks before and up to 6 months after a 3-day course of 3rd-generation cephalosporins. Our results show that gut phages rarely encode antibiotic resistance genes and are mostly temperate, including many phage plasmids. Furthermore, phage populations remain individual-specific even after microbiome perturbation. Yet, we report a 20% decline in phage diversity the day after treatment, alongside blooms of a few, mostly virulent, phages. We suggest that some of these phages contribute to the recovery of bacterial diversity via “kill-the-winner” dynamics. This is supported by (temporarily) dominant phages targeting Parabacteroides distasonis, a bacterium that thrives post-treatment only in the absence of these phages. Our findings suggest gut phages are crucial to the microbiome response to antibiotics, aiding the restoration of balance and diversity.
Recommended citation: E. Pfeifer, C. d’Humières, Q. Lamy-Besnier, F.P. Oñate, R. Denise, S. Dion, B. Condamine, M. Touchon, L. Ma, C. Burdet, F. Mentré, E. Denamur, E.P.C. Rocha, Antibiotic perturbation of the human gut phageome preserves its individuality and promotes blooms of virulent phages. Cell Reports, 44(8):116020, 2025. https://pubmed.ncbi.nlm.nih.gov/40694475/
Published in mBio, 2025
Bacteriophages (phages) are increasingly explored as therapeutic agents against bacterial infections. However, most studies focus on in vitro systems, offering limited insight into the complexity of such dynamics in physiological contexts. Here, we use the mouse-restricted enteric pathogen Citrobacter rodentium (CR), a well-established model for human enteropathogenic and enterohemorrhagic Escherichia coli (EPEC and EHEC) infections, to investigate phage-pathogen interactions in a murine model with a complex microbiota. We isolate and characterize Eifel2, a novel lytic phage infecting CR, and generate anti-phage-specific antibodies that enable the visualization of phage infections in vitro. In a murine model of CR infection, oral administration of Eifel2 led to robust phage replication in the gut without reducing the bacterial burden or infection-associated inflammation, confirming the establishment of a stable co-existence in the gut. Despite the emergence of a sub-population of phage-resistant CR mutants in vivo, they did not undergo clonal expansion, indicating that additional selective pressures impaired their widespread dissemination in the gut. Together, our findings demonstrate that imaging approaches can capture key infection stages in vitro, although in vivo models are essential for capturing the complexity of phage-bacteria interactions. This work highlights the importance of studying phage therapy in host-pathogen contexts that include a normal microbiota and a suitable host environment, where dynamic co-existence rather than eradication may define therapeutic outcomes.
Recommended citation: A. Peters, H. Shareefdeen, J. Sanchez-Garrido, E. J. Cohen, R. Denise, J. L. C. Wong, M. Beeby, C. Hill, G. Frankel, Stable co-existence of Citrobacter rodentium with a lytic bacteriophage during in vivo murine infection. mBio, 2025. https://pubmed.ncbi.nlm.nih.gov/41460104/
Published:
Published:
Published:
Published:
Poster presented at SMBE 2024.
Published:
Poster presented at ISBA 2025.
Undergraduate course, Pierre and Marie Curie University, Sorbonne University, 2016
Course: LV103 / LU1SV002
Undergraduate course, Pierre and Marie Curie University, Sorbonne University, 2018
Course: LV101 / LV103 / LU1SV001
Undergraduate course, Pierre and Marie Curie University, Sorbonne University, 2018
Course: LU2SV385
Summer school, Department of Archaeogenetics, Max Planck Institute for Evolutionary Anthropology, 2024